






The Spectra Medical sodium chloride flush recall covers specific 0.9% Sodium Chloride Injection USP 10 mL ampules that FDA says should be removed from stock and use because manufacturing and sterility concerns could expose patients to microorganisms, endotoxins or foreign matter. FDA’s September 18, 2026 public communication identifies the action as its most serious type of medical-device recall. However, Spectra had already initiated the underlying recall on July 6, so September 18 should not be described as the original recall date.
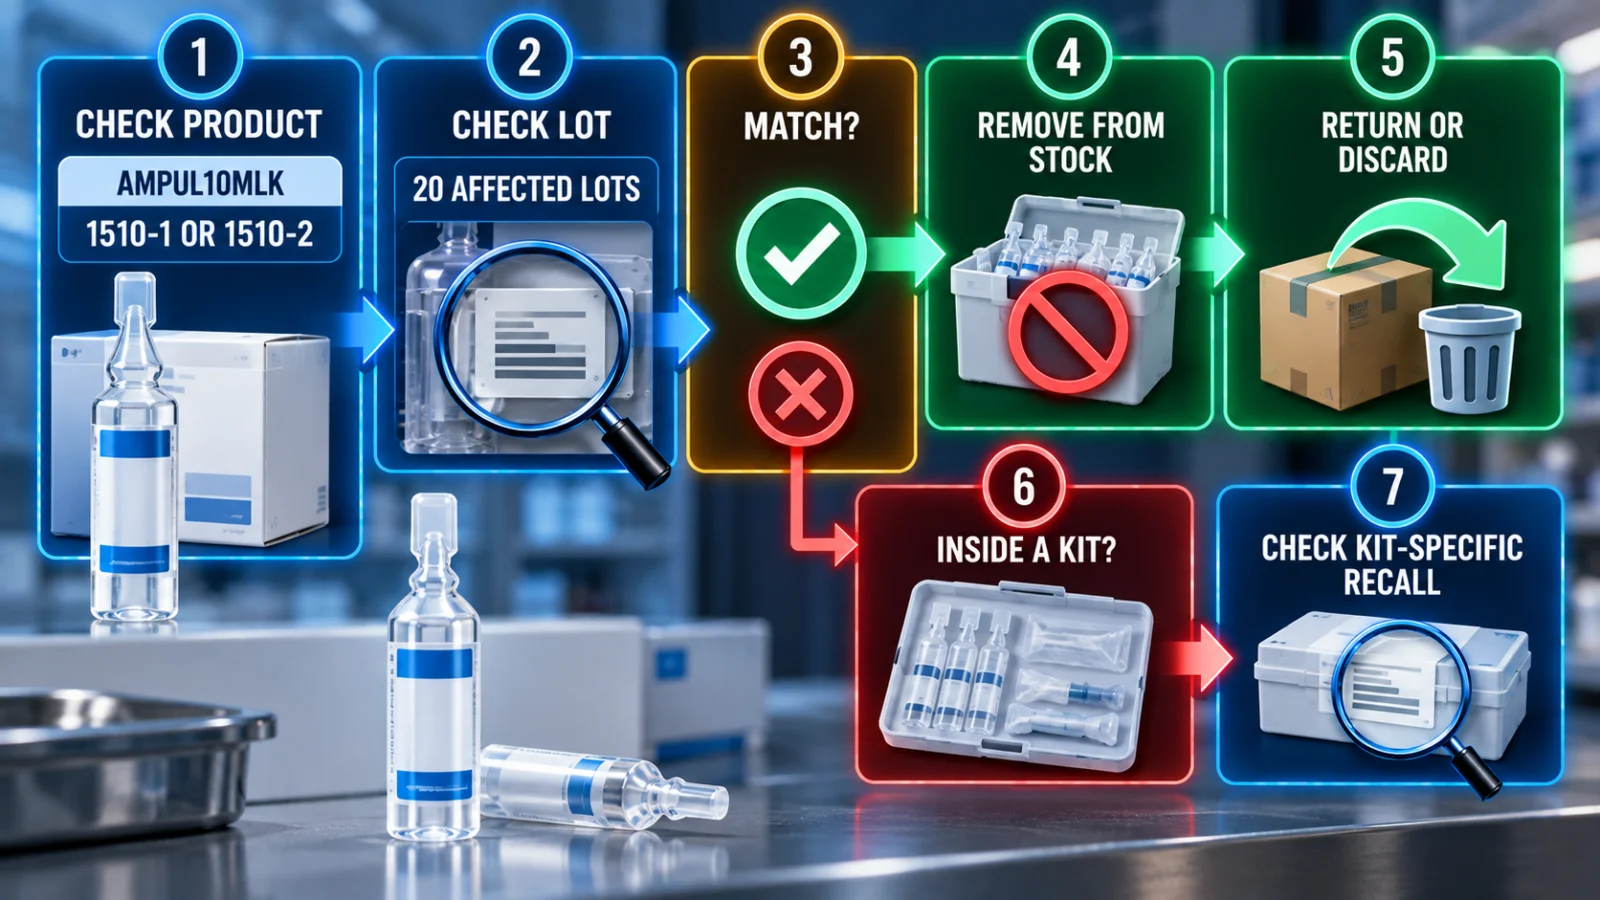
What facilities should do: check the product identity, catalog or product code and lot number. Affected product is identified as Sodium Chloride Injection 0.9% USP, 10 mL ampule, model/catalog AMPUL10MLK, including product codes 1510-1 and 1510-2. If the lot matches FDA’s affected population, remove unused ampules from stock and use and follow the current return or disposal instructions. If the ampule is inside a convenience kit, also check the kit-specific recall instructions before deciding what happens to the remaining kit components.
Which Spectra sodium chloride flush lots are affected?
FDA’s enforcement record lists 20 affected lot codes:
- AB3005
- AB3006
- AB3007
- AB3008
- AB3009
- AB3010
- AB4001
- AB4002
- AB4003
- AB4004
- AB4005
- AB4006
- AB5001
- AB5002
- AB5003
- AB5004
- AB5005
- AB5006
- AB5007
- AB5008
Facilities should match the exact product and lot information rather than treating every sodium chloride flush or every Spectra product as recalled.
How to verify affected inventory
Check for Sodium Chloride Injection 0.9% USP in a 10 mL ampule and identify catalog AMPUL10MLK or product code 1510-1 or 1510-2.
Compare the lot code with FDA’s 20 affected AB-series lots.
Affected unused ampules should be removed from stock and use immediately.
Use the current recall instructions for returning or discarding affected unused product and for notifying downstream customers when applicable.
If an affected ampule is packaged inside a medical kit, verify the kit manufacturer’s own recall or correction instructions before disposing of the entire kit.
Why is FDA treating this as a serious recall?
FDA says manufacturing concerns could compromise product quality or sterility. If sterility is compromised, patients could be exposed to microorganisms, endotoxins or foreign matter. FDA describes potential consequences including systemic inflammatory responses, infections, micro-clots and organ injury or failure. These are potential outcomes from exposure to affected product, not proof that every recalled ampule is contaminated or that every exposed patient will experience harm.
Have serious injuries or deaths been reported?
FDA’s September communication says Spectra had reported no serious injuries or deaths associated with the issue as of July 6, 2026. That statement is date-specific and should not be interpreted as a permanent adverse-event count. Later FDA or manufacturer updates could change the record.
Why July 6 and September 18 are both important
Spectra initiated the underlying recall on July 6. FDA later published the current public-facing recall communication on September 18 and highlighted the action as the most serious type of medical-device recall. The September FDA communication therefore increases current awareness and urgency, but it did not create the original recall.
What if the ampule is inside a convenience kit?
FDA records show that Spectra distributed affected product to medical-device kit packers, and downstream kit manufacturers have issued separate actions involving Spectra-recalled sodium chloride ampules. The correct response may depend on the kit-specific notice. In some downstream actions, the affected ampule is removed while other kit components can remain usable. Do not assume that every kit containing a Spectra ampule has identical instructions.
Where was the product distributed?
FDA’s enforcement record lists direct distribution to medical-device kit packers in California, Florida, North Carolina, Oklahoma, Texas and Virginia. That direct-distribution list does not necessarily define the final end-user geography after downstream kit distribution, so facilities outside those states should not use geography alone to rule out affected inventory.
Verification and limitations
The central recall classification, product identifiers, 20 lot codes, hazard pathway and removal instructions were checked against FDA’s September 18 public recall communication and FDA enforcement record Z-2873-2026, with Spectra’s July 6 company notice used to reconcile the original recall timeline. The total quantity in commerce, ultimate downstream geographic reach, post-July-6 adverse-event status and future recall termination date remain unresolved and may change.
Bottom line: facilities should verify the exact Spectra product and lot rather than generalizing this recall to all saline flushes. Matching affected inventory should be removed from stock and use, with return or disposal handled under the current recall instructions and convenience-kit stock checked against its own downstream notice when applicable.